Unconfigured Ad

Collapse
X
 
  • Time
  • Show
Clear All
new posts
  • rndouglas
    Member
    • Jun 2012
    • 23

    #1

    Bowtie mapping with more than 3 mismatches?

    I'm trying to map some sRNA reads to specific loci of interest while allowing up to 3 mismatches using -v 3.

    For some of my target loci, this seems to work well. All of the mapped reads have no more than 3 mismatches.

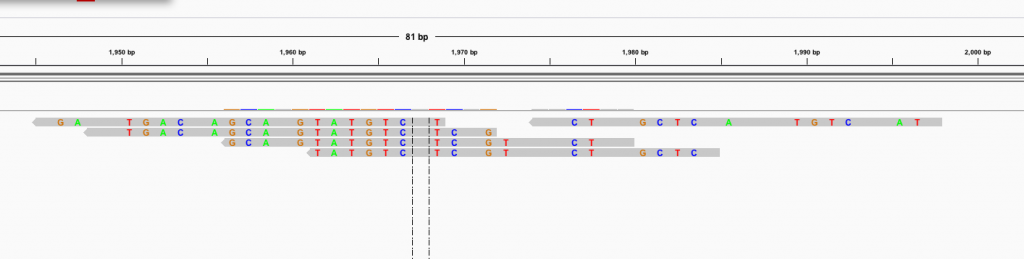
    However, when I use the exact same code, changing only the bowtie files for a new locus, I'll get reads with up to 20 mismatches (and the sRNA read is only 24nt long!).

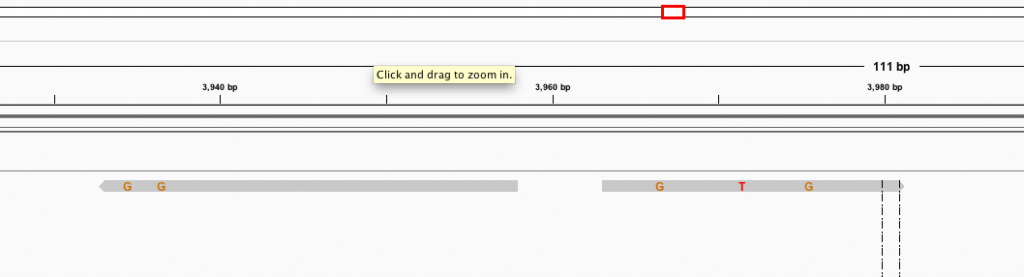
    I've attached two screenshots from IGV showing the same sRNA sample mapped with Bowtie against two different loci. All of the nucleotides shown are mismatches.

    How I expect it to always look:

    How it usually turns out:

    The code I'm using is as follows:

    Code:
    bowtie -f -v 3 -S ~/CRM2 ~/sRNA.fa > ~/output.sam
    Oddly, I also have the exact same issue (too many mismatches) when I use this code:
    Code:
    bowtie -f -n 0 -l 25 -S ~/CRM2 ~/sRNA.fa > ~/output.sam
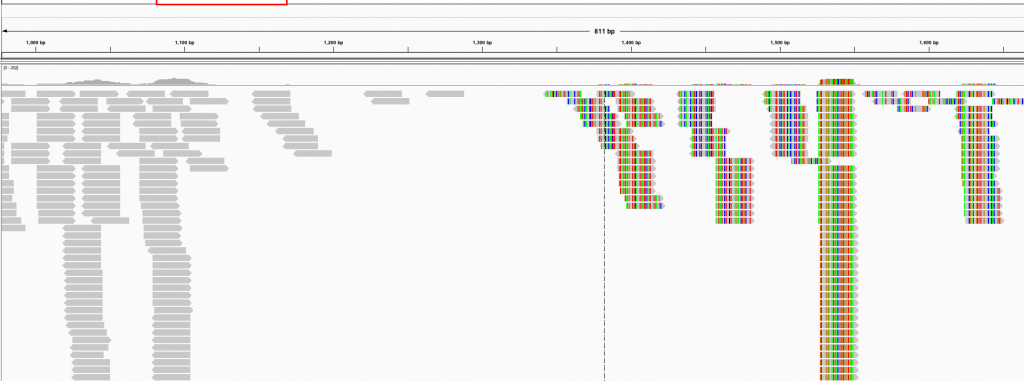
    Am I doing something wrong? Has anyone else seen this?
  • rndouglas
    Member
    • Jun 2012
    • 23

    #2
    Cranking up the frustration a notch....

    This week I discovered the same situation (up to 20 mismatches on a 24 nt "mapped" read) occurs when I set -v to 0, supposedly not allowing for mismatches.


    This happens with ~4-5 loci I'm interested in mapping to, but several other loci of similar length do not show this problem at all.

    Comment

    • Nepher
      Junior Member
      • Jan 2013
      • 4

      #3
      Have the same problem

      Hi all,

      since this thread has no answer and that I came across exact the same issue I wanted to know if someone knows how this type of alignment is possible. Where is the mistake ?
      I also have 24 mismatches on sequences that were mapped with NO mismatch option...

      If you could help I would be very grateful

      Thanks a lot

      Comment

      • rndouglas
        Member
        • Jun 2012
        • 23

        #4
        For me, it was an issue with IGV. When I looked more closely IGV was coloring nucleotides as being mismatched, but they really were not mismatched. I never found a workaround/explanation, so I just had to ignore all of the claimed mismatches.

        Bowtie was mapping properly with zero mismatches allowed.

        Comment

        • Nepher
          Junior Member
          • Jan 2013
          • 4

          #5
          Thanks for your answer.

          I came across a solution this morning. After checking lots of stuff I looked again at my reference sequence and I saw that there were special characters at the beginning of the sequence :

          >name
          ??????ATCGTGCTA

          I simply erased them and IGV removed the most part of mismatches (it left those from the mapping). I think that the mapping coordinates where simply shifted/reference sequence.

          Hope this will help other users with IGV problems

          Comment

          Latest Articles

          Collapse

          • SEQadmin2
            Multiomic Liquid Biopsy: What DNA/RNA Panels Reveal That DNA Alone Can’t
            by SEQadmin2


            Tumors contain complex mixtures of cell populations that evolve in response to treatment and host-specific factors. Tissue biopsies provide only static snapshots of a tumor’s state. But what if we could follow that evolution over time using a minimally invasive technique?

            Liquid biopsies can detect tumor-derived material circulating in the bloodstream, providing a way to study treatment response, detect emerging resistance, and discover new biomarkers. Circulating
            ...
            10-14-2026, 09:55 AM
          • SEQadmin2
            New Genomics Technologies Take Aim at Long-Standing Limits
            by SEQadmin2


            Researchers using sequencing and genomics tools often have to make trade-offs. They can choose between speed or scale, short reads or long-range information, or targeted panels or a view of the whole transcriptome. New technologies that have been released this year are built to address those tough choices.

            We asked six companies the same four questions to learn about their latest products. The new technologies bring a lot to the table, including rethinking sequencing
            ...
            09-28-2026, 10:25 AM

          ad_right_rmr

          Collapse

          News

          Collapse

          Topics Statistics Last Post
          Started by SEQadmin2, 10-22-2026, 11:13 AM
          0 responses
          24 views
          0 reactions
          Last Post SEQadmin2  
          Started by SEQadmin2, 10-22-2026, 10:48 AM
          0 responses
          20 views
          0 reactions
          Last Post SEQadmin2  
          Started by SEQadmin2, 10-15-2026, 10:52 AM
          0 responses
          36 views
          0 reactions
          Last Post SEQadmin2  
          Started by SEQadmin2, 10-14-2026, 05:39 AM
          0 responses
          42 views
          0 reactions
          Last Post SEQadmin2  
          Working...