Unconfigured Ad

Collapse
X
 
  • Time
  • Show
Clear All
new posts
  • pbabb926
    Junior Member
    • Oct 2012
    • 3

    #1

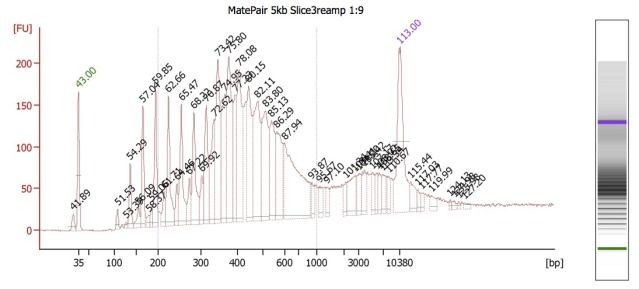
    Odd "rooster-comb" BioAnalyzer profile for Mate Pair library

    Greetings all!

    Many thanks to everyone who has contributed to this very helpful and informative forum. This is my first post, so I figured I'd make things a bit interesting with a strange scenario I've recently encountered...

    I'm currently undertaking library preparation using a bit of a hybrid of Illumina's TruSeq and MatePair kits and protocols. The goal is to generate three libraries of different insert sizes - roughly 300 bp, 3kb and 5kb. These different insert-size-libraries are then pooled at a specific ratio and run across multiple lanes on a HiSeq flow cell.

    Because these different insert-size-libraries are pooled, they require indexed adapters to sort out their library (and thus, insert size) of origin. This is where the hybrid protocol comes in: the Mate Pair kit does not have indexed adapters, so we've integrated TruSeq adapters into this protocol. So far, this worked well for creating the 3kb libraries, but for some reason, this did NOT work well for the 5kb libraries. BioAnalyzer results (and agarose gels) showed the presence of many, regularly spaced, fragment bands.


    We believe that these are primer dimers/concatemers of some kind, but are puzzled on why they would appear for the 5kb libraries but not the 3kb ones. Both Mate Pair library types were set up at the same time, using the same DNA, and utilizing nearly identical workflow procedures.

    Has anyone out there in Next-Generation-Sequencing-land experienced similar results??

    Many thanks!
    Paul
  • pmiguel
    Senior Member
    • Aug 2008
    • 2328

    #2
    Every once in a while we see a library like this. "rooster comb", eh?

    Anyway, I have no real answers, but can add a data point and speculate.

    Constructed a set of TruSeq RNA libraries using normal TruSeq RNA (would be called "LT", these days). Except these are going to be run on the MiSeq using 2 x 250. So wanted the inserts longer than normal. I won't go into the gory details, but we used 2 minute for the divalent cation/heat framentation step instead of the standard 8 minutes. The RNA was somewhat low in mRNA, I think, so we were limiting for cDNA at the ligation step.

    After ligation + the initial 1:1 ampure clean-up the second size selection was done. 2 ampure fractions were taken. One larger fraction at 1:0.6 sample:ampure then, the supernatant from that fraction had another full volume of ampure added to it. Which probably pulled down nearly everything. Each sample was subjected to 15 cycles of enrichment PCR, as is specified in the protocol.

    The larger fraction looked fairly normal:


    The lower molecular weight fraction however has "combed-out":


    Notable to me is the enormous adapter dimer peak (around 120 bp) in the lower molecular weight fraction. So my tendency would be:

    (1) Blame everything on the low insert:adapter ratio causing high adapter-dimer contamination of the pre-amp library.
    (2) Further speculate that the second ampure cut largely, or maybe completely removed that adapter-dimer resulting in small or non-existent spines.
    (3) But the rescue of the low-molecular weight fraction resulted in a high concentration of that adapter dimer -- and this dimer caused the spines somehow.
    (4) The spines probably result from the adapter dimers annealing to full length library products -- possibly being extended by the PCR polymerase into "insert+adapter" products. Additional additions could occur during subsequent cycles of PCR.

    I should point out that this hypothesis is not particularly supported by the data at hand. 3 other libraries produced at the same time did not grow spines, despite having the same enormous adapter-dimer peaks in them.

    So, maybe the adapter dimer is not a contributor to this effect. At best I guess it might be necessary but not sufficient.

    But when I do see spines, they are in libraries produced with limiting mRNA. So, in your case I would tend to attribute the phenomenon to low insert DNA amounts, possibly the result of poor conversion of first fragmentation DNA into circular DNA in your 5 kb reaction.

    --
    Phillip
    Last edited by pmiguel; 10-10-2012, 07:03 AM.

    Comment

    • pbabb926
      Junior Member
      • Oct 2012
      • 3

      #3
      Thanks so much for adding data and suggestions to our query, Phillip!

      Your hypotheses seem quite logical, and in line with our own internal guesswork.
      We suspect that the amount of material moving downstream in our 5kb pipeline was lower in quantity than our other library constructs, and that we didn't meet some crucial ratio-based threshold in the adapter ligation and/or subsequent purification and enrichment steps.

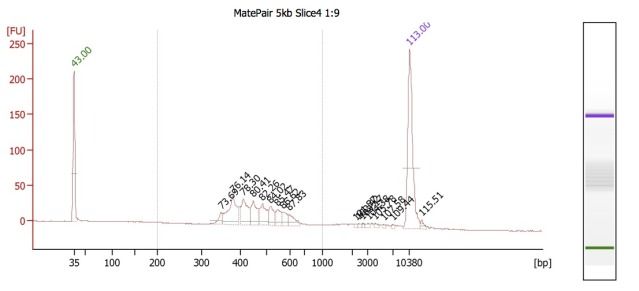
      We also experimented with re-amplifing the biotinylated fragments off of the streptavidin beads in the Mate Pair protocol, but that seemed to exacerbate our initial problems and left us with a similar BioA profile as the second picture you posted above:


      We're going to be reconstructing these larger insert libraries again in short order, so we will have an opportunity to implement a few changes to our protocol in an effort to cut down these crazy adapter-dimers and concatamers.

      If anyone has any specific recommendations, we're all ears!


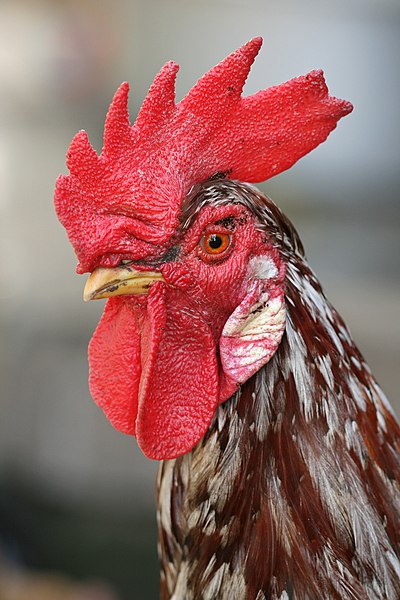
      Here is a real rooster-comb for comparison

      Comment

      • pmiguel
        Senior Member
        • Aug 2008
        • 2328

        #4
        For some reason your post brought to mind the original Gremlins movie. At least some of the "bad" gremlins had swordfish or dimetrodon spined fin on their heads. Also the conversion of the fuzzy Gizmo-like gremlins to their malevolent form has a certain resonance with a spined library distribution on an agilent chip.

        --
        Phillip

        Comment

        Latest Articles

        Collapse

        • SEQadmin2
          Multiomic Liquid Biopsy: What DNA/RNA Panels Reveal That DNA Alone Can’t
          by SEQadmin2


          Tumors contain complex mixtures of cell populations that evolve in response to treatment and host-specific factors. Tissue biopsies provide only static snapshots of a tumor’s state. But what if we could follow that evolution over time using a minimally invasive technique?

          Liquid biopsies can detect tumor-derived material circulating in the bloodstream, providing a way to study treatment response, detect emerging resistance, and discover new biomarkers. Circulating
          ...
          10-14-2026, 09:55 AM
        • SEQadmin2
          New Genomics Technologies Take Aim at Long-Standing Limits
          by SEQadmin2


          Researchers using sequencing and genomics tools often have to make trade-offs. They can choose between speed or scale, short reads or long-range information, or targeted panels or a view of the whole transcriptome. New technologies that have been released this year are built to address those tough choices.

          We asked six companies the same four questions to learn about their latest products. The new technologies bring a lot to the table, including rethinking sequencing
          ...
          09-28-2026, 10:25 AM

        ad_right_rmr

        Collapse

        News

        Collapse

        Topics Statistics Last Post
        Started by SEQadmin2, 10-22-2026, 11:13 AM
        0 responses
        23 views
        0 reactions
        Last Post SEQadmin2  
        Started by SEQadmin2, 10-22-2026, 10:48 AM
        0 responses
        20 views
        0 reactions
        Last Post SEQadmin2  
        Started by SEQadmin2, 10-15-2026, 10:52 AM
        0 responses
        36 views
        0 reactions
        Last Post SEQadmin2  
        Started by SEQadmin2, 10-14-2026, 05:39 AM
        0 responses
        42 views
        0 reactions
        Last Post SEQadmin2  
        Working...