Unconfigured Ad

Collapse
X
 
  • Time
  • Show
Clear All
new posts
  • frymor
    Senior Member
    • May 2010
    • 151

    #1

    trimming in tophat

    Hi all,

    I am trying to analyse my PE Illumina data using tophat.

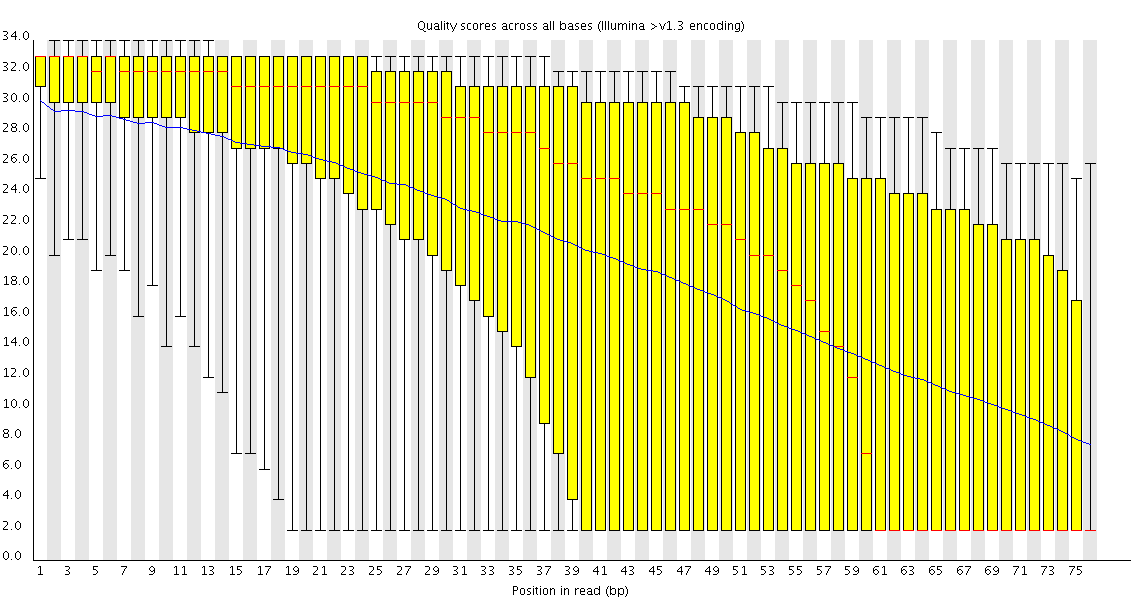
    At first I run fastqc. Checking the raw data, I discovered at the beginnings (and presumably at the ends) of my reads I have some containments from the adapters of the sequencing.
    I run bowtie first on both the full length and trimmed sequences and got better results with the trimmed sequences.

    Do I need to trim the data before running tophat?

    Does someone know how to do it? do I need to convert my trimmed sam files (bowtie output) back into fastq files?

    Thanks for any help
    Assa
  • dnusol
    Senior Member
    • Jul 2009
    • 136

    #2
    Hi, I found this useful page about this issue.


    HTH

    Dave

    Comment

    • frymor
      Senior Member
      • May 2010
      • 151

      #3
      Thanks for the tip.
      It is a good page with summaries about the different images of the fastqc software, a thing a lot of people were looking for in a different thread.

      BTW, can anyone tell me of a good way to remove the duplicates reads from the equation.
      running the fastqc program I get a lot of duplicated reads (see attachment).

      As I am looking for differentially regulated genes I am not sure whether I should exclude the duplicated reads or not, but I would like to try ans see what I get when doing so.

      Q: can anyone tell me how to filter duplicated genes from the sam files or before the bowtie run from the fastq files?

      Q: Is it the right way when going for differential expression also to exclude the duplications? or do I need to keep them?

      Thanks

      Assa
      Attached Files

      Comment

      • jp.
        Senior Member
        • Jul 2013
        • 142

        #4
        did you get the answer ?
        would like to share it here
        thank you

        Originally posted by frymor View Post
        Thanks for the tip.
        It is a good page with summaries about the different images of the fastqc software, a thing a lot of people were looking for in a different thread.

        BTW, can anyone tell me of a good way to remove the duplicates reads from the equation.
        running the fastqc program I get a lot of duplicated reads (see attachment).

        As I am looking for differentially regulated genes I am not sure whether I should exclude the duplicated reads or not, but I would like to try ans see what I get when doing so.

        Q: can anyone tell me how to filter duplicated genes from the sam files or before the bowtie run from the fastq files?

        Q: Is it the right way when going for differential expression also to exclude the duplications? or do I need to keep them?

        Thanks

        Assa

        Comment

        • frymor
          Senior Member
          • May 2010
          • 151

          #5
          No I didn't get any response for the questions I posted.

          I am not sure though how important is the duplication rate in this step. I'm using tophat2 with the option to exclude all duplicated reads, so I am not worried about the duplication in the original fastq file.

          I hope I am thinking in the right direction.

          Comment

          • anamika
            Junior Member
            • Apr 2013
            • 5

            #6
            Sangenix

            SangeniX: A comprehensive, automated, scalable and user friendly NGS data analysis suite

            Sangenix Has module for duplication removal.

            Give it a try : http://www.sangenix.com/

            Comment

            • frymor
              Senior Member
              • May 2010
              • 151

              #7
              let me know again, when it is a freeware

              Comment

              • vineet jha
                Junior Member
                • Feb 2013
                • 4

                #8
                Sangenix

                Beta Version is available. you can contact to us via contact page in http://www.sangenix.com/contactus.aspx

                Comment

                • dpryan
                  Devon Ryan
                  • Jul 2011
                  • 3478

                  #9
                  Removing the duplicates could be done with the samtools rmdup command (you could alternatively use markDuplicates from picard). This is generally not needed for RNAseq, since a certain amount of duplication would be both expected and desired for highly expressed genes (i.e., many/most of these probably aren't PCR duplicates).

                  Comment

                  Latest Articles

                  Collapse

                  • SEQadmin2
                    Multiomic Liquid Biopsy: What DNA/RNA Panels Reveal That DNA Alone Can’t
                    by SEQadmin2


                    Tumors contain complex mixtures of cell populations that evolve in response to treatment and host-specific factors. Tissue biopsies provide only static snapshots of a tumor’s state. But what if we could follow that evolution over time using a minimally invasive technique?

                    Liquid biopsies can detect tumor-derived material circulating in the bloodstream, providing a way to study treatment response, detect emerging resistance, and discover new biomarkers. Circulating
                    ...
                    10-14-2026, 09:55 AM
                  • SEQadmin2
                    New Genomics Technologies Take Aim at Long-Standing Limits
                    by SEQadmin2


                    Researchers using sequencing and genomics tools often have to make trade-offs. They can choose between speed or scale, short reads or long-range information, or targeted panels or a view of the whole transcriptome. New technologies that have been released this year are built to address those tough choices.

                    We asked six companies the same four questions to learn about their latest products. The new technologies bring a lot to the table, including rethinking sequencing
                    ...
                    09-28-2026, 10:25 AM

                  ad_right_rmr

                  Collapse

                  News

                  Collapse

                  Topics Statistics Last Post
                  Started by SEQadmin2, 10-22-2026, 11:13 AM
                  0 responses
                  57 views
                  0 reactions
                  Last Post SEQadmin2  
                  Started by SEQadmin2, 10-22-2026, 10:48 AM
                  0 responses
                  41 views
                  0 reactions
                  Last Post SEQadmin2  
                  Started by SEQadmin2, 10-15-2026, 10:52 AM
                  0 responses
                  46 views
                  0 reactions
                  Last Post SEQadmin2  
                  Started by SEQadmin2, 10-14-2026, 05:39 AM
                  0 responses
                  46 views
                  0 reactions
                  Last Post SEQadmin2  
                  Working...